
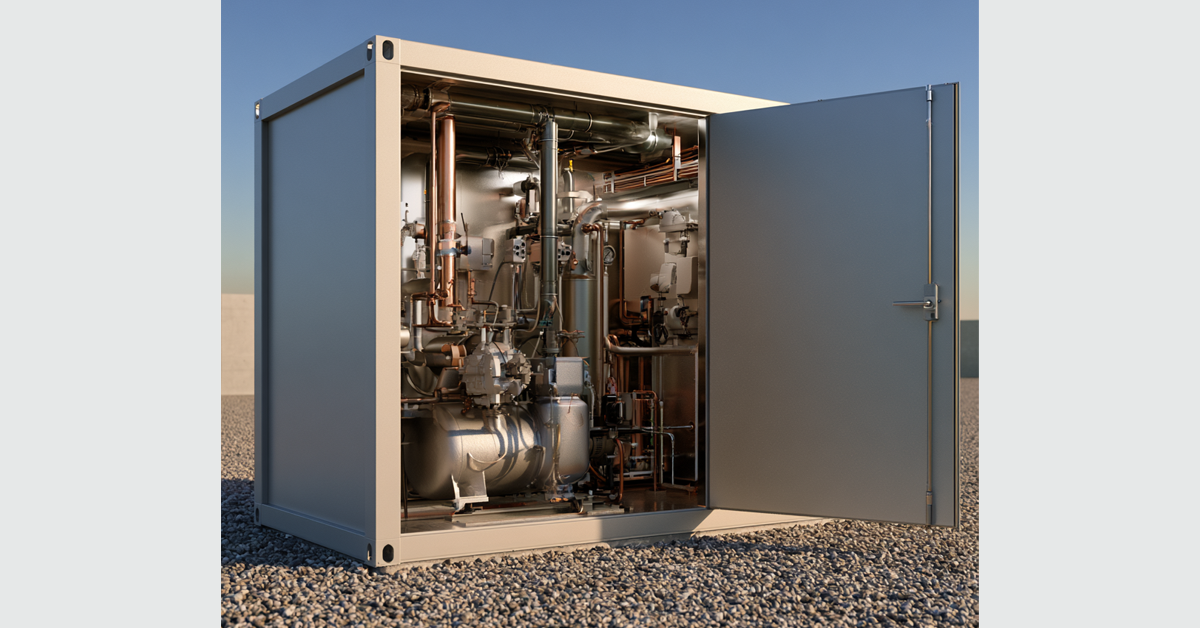
Factory-assembled packaged mechanical systems are increasingly specified for pharmaceutical utility applications. The advantages of factory assembly, compressed installation timelines, reduced field coordination complexity, documented factory testing, and single-point utility connections, are particularly valuable in pharmaceutical facilities where the installation must be coordinated around ongoing manufacturing operations and where the qualification process requires documented evidence of how the system was built and tested.
The regulatory environment that pharmaceutical utility systems operate in places requirements on the equipment and the documentation that standard commercial packaged system specifications do not address. Engineers and facility managers specifying packaged systems for pharmaceutical utility applications benefit from understanding what those requirements mean before the supplier is selected and the specification is written.
What cGMP means in the context of packaged mechanical systems
Current Good Manufacturing Practice regulations, enforced by the FDA under 21 CFR Part 211 for pharmaceutical manufacturing, establish requirements for equipment design, construction, and maintenance in regulated facilities. The regulations require that equipment used in pharmaceutical manufacturing be of appropriate design and construction, be installed and maintained properly, and be capable of performing its intended function consistently and reproducibly.
For packaged mechanical systems serving pharmaceutical utilities, cGMP compliance is not a certification that a supplier receives. It is a set of requirements that the equipment and the documentation need to satisfy in the context of the specific facility and the specific utility system. A packaged pump skid serving a purified water circulation system needs to be designed, constructed, documented, and maintained in a way that the facility’s quality system can verify. What that means specifically depends on the utility system, the facility’s quality standards, and the regulatory requirements that apply to the manufacturing operations the utility supports.
The practical implication for engineers specifying packaged systems is that the supplier’s documentation capability matters as much as the system’s mechanical performance. A packaged system that performs correctly but cannot be documented to the facility’s qualification standards cannot be fully validated, which limits its regulatory acceptance regardless of how well it works.
Material traceability
Material traceability in a cGMP context means that the materials used in the construction of the packaged system can be documented from the mill certificate for the raw material through the fabrication process to the finished component in the system. For wetted components in pharmaceutical utility systems, this traceability is part of the installation qualification documentation that demonstrates the system was built with the specified materials.
Engineers specifying packaged systems for pharmaceutical utility applications should confirm with the supplier what material documentation they can provide, what the format of that documentation is, and how it will be organized in the documentation package that accompanies the system. A supplier who provides mill certificates for the major pressure-containing components but cannot provide documentation for the elastomers, the fasteners, and the instrumentation connections leaves gaps in the material traceability record that the IQ process will need to address.
The material traceability requirement also extends to any changes made to the system during fabrication. If a specified component is unavailable and a substitution is made, the substitution needs to be documented, reviewed against the specification requirements, and approved before the component is installed. A change that is made without documentation creates a gap in the traceability record that may not be discovered until the IQ review, at which point the system may need to be partially disassembled to confirm what was actually installed.
Weld documentation
For packaged systems that include welded pressure piping or welded vessels in pharmaceutical utility applications, weld documentation is a component of the IQ package. Weld documentation typically includes the welder qualifications, the weld procedure specifications, and the weld inspection records for each weld in the system.
Orbital welding is commonly used in pharmaceutical utility piping for its consistency and reproducibility. An orbital weld made with a qualified procedure by a qualified operator on a properly prepared joint produces a consistent, documented result that manual welding cannot match for uniformity. For applications where weld quality and documentation are part of the qualification requirement, confirming that the supplier uses orbital welding procedures and maintains weld logs as part of their standard fabrication documentation is an important specification question.
For systems where weld inspection is required, the inspection method and the acceptance criteria need to be defined in the specification. Visual inspection, borescope inspection, and in some applications radiographic inspection of completed welds are used depending on the system pressure, the fluid service, and the facility’s quality requirements. The inspection records need to be organized as part of the IQ documentation package.
Factory testing documentation for qualification
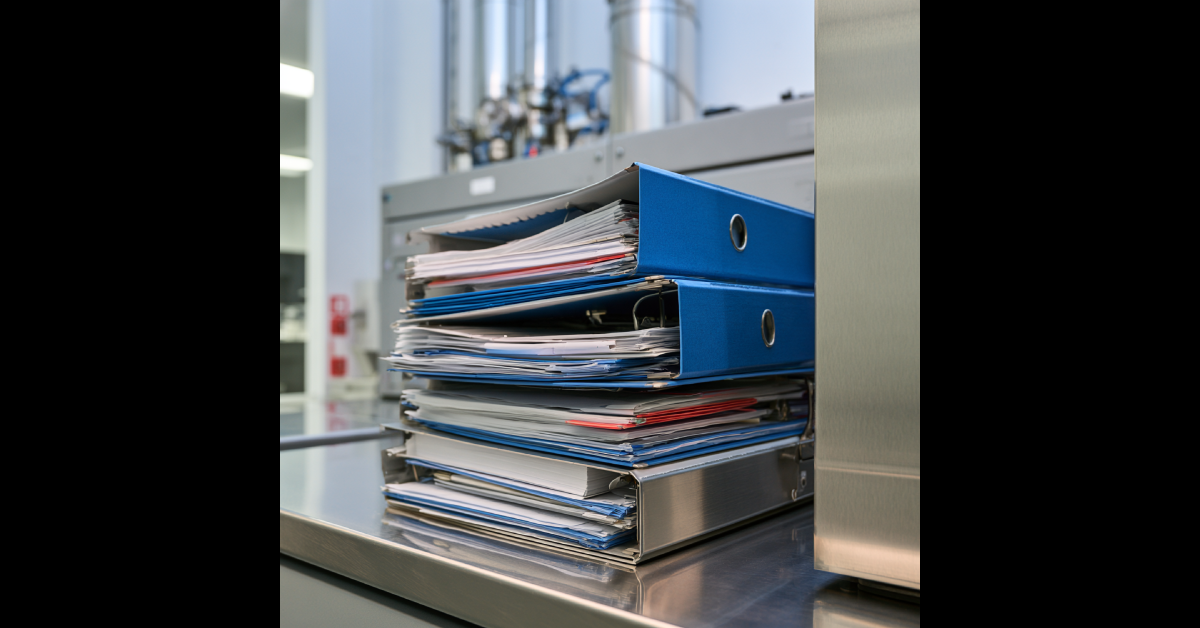
The factory testing documentation for a packaged system destined for a pharmaceutical utility application needs to be organized to support the IQ process, not just to confirm that the system was tested. The IQ reviewer needs to be able to trace from the specified test requirements to the documented test results for each requirement.
A factory test report that records the results of pressure testing, alignment verification, controls testing, and functional run testing in a format organized by test requirement, with the specified acceptance criteria and the measured results clearly recorded, is significantly more useful for IQ purposes than a summary test report that confirms testing was performed without the detail required to complete the IQ review.
Engineers specifying packaged systems for pharmaceutical utility applications should request a sample factory test report format from the supplier before the purchase order is written. The sample report shows what documentation the supplier produces as standard practice and what additional documentation may need to be requested. If the supplier’s standard test documentation does not support the facility’s IQ requirements, that is better discovered before fabrication than after the system is complete and on site.
Questions to ask packaged system suppliers
Before a purchase order is placed for a packaged mechanical system destined for a pharmaceutical utility application, engineers should confirm the following with the supplier. What material documentation does the supplier provide for wetted components, and does it include mill certificates, elastomer specifications, and instrumentation certifications? How does the supplier document material substitutions made during fabrication? Does the supplier maintain weld logs and welder qualification records, and are orbital welding procedures used for pharmaceutical utility piping? What does the standard factory test report look like, and can the supplier provide a sample? How is the complete documentation package organized for delivery with the system?
The answers to these questions do not guarantee that the packaged system will satisfy the facility’s qualification requirements, because those requirements are specific to the facility and the utility system. But they establish whether the supplier has the documentation capability and the quality discipline that pharmaceutical utility applications require, before the commitment to a specific supplier is made.
For pharmaceutical utility applications where the packaged system includes GP Energy Products steam system equipment, the GP Energy team handles the steam system specification and can coordinate with FabPro on system integration. Visit gpenergyproducts.com for more. For applications where Merion Pump Company equipment is part of the package, the Merion team handles pump specification for pharmaceutical utility applications. Visit merionpump.com for more. For heat exchanger coils in pharmaceutical utility packaged systems, the HX Coils team handles coil specification for regulated environments. Visit hxcoils.com for more.
References
1. FDA. 21 CFR Part 211. Current Good Manufacturing Practice for Finished Pharmaceuticals. Governs equipment design, construction, and documentation requirements in FDA-regulated manufacturing facilities. fda.gov
2. ISPE. Good Practice Guide: Commissioning and Qualification. Covers IQ/OQ/PQ requirements for utility systems in pharmaceutical manufacturing facilities. ispe.org
3. ASME BPE. Bioprocessing Equipment Standard. Covers design, materials, and fabrication requirements for equipment used in pharmaceutical and bioprocessing applications. asme.org
4. ASTM A270. Standard Specification for Seamless and Welded Austenitic Stainless Steel Sanitary Tubing. Covers material and weld requirements for stainless steel components in pharmaceutical utility applications. astm.org


Leave A Comment